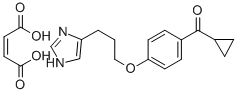
Ciproxifan is a histamine H3 receptor antagonist (Ki = 0.5 nM for inhibition of histamine release in rat synaptosomal preparations). It is selective for histamine H3 over histamine H1 and H2, M3 muscarinic, and α1D- and β1-adrenergic receptors (Kis = 3.15-25 μM), as well as the serotonin (5-HT) receptor subtypes 5-HT1B, 5-HT2A, 5-HT3, and 5-HT4 (Kis = 1.58-15 μM) in radioligand binding assays. Ciproxifan (3-300 nM) inhibits relaxation in precontracted isolated guinea pig ileal longitudinal muscle induced by R-(–)-α-methylhistamine . In vivo, ciproxifan reduces R-(–)-α-methylhistamine-induced water consumption in rats (ED50 = 0.09 mg/kg). It increases wake episode duration and latency to fall asleep in rats when administered at a dose of 2 mg/kg.
Ciproxifan is a potent histamine H3 receptor antagonist which affects histamine levels in the brain. It may be used in treatments for sleep and neurological disorders.
Potent, selective H3 histamine receptor antagonist.
Ciproxifan (1 mg/kg; a single p.o.) increases the t-MeHA level in mouse brain, with an ED50 of 0.14 mg/kg[1].
Ciproxifan (3 mg/kg, i.p.) improves the accuracy of responding in the five-choice task in rats only when the stimulus duration is 0.25 sec instead of 0.50 sec[1].
Ciproxifan (0.15-2 mg/kg; p.o.) induces marked signs of neocortical electroencephalogram activation manifested by enhanced fast-rhythms density and an almost total waking state in cats[1].
Ciproxifan (1 mg/kg; a single i.v.) decreases the H3-receptor ligand concentration in serum in mice, with the half-lives (t1/2) of 13 and 87 min for the distribution and elimination phases in mice, respectively[1].
Ciproxifan (1 mg/kg; a single p.o.) exhibits oral bioavailability (F=62%) and maximal concentration (Cmax=420 nM) in mice[1].
H3 receptor: 9.2 nM (IC50)
[1] X LIGNEAU. Neurochemical and behavioral effects of ciproxifan, a potent histamine H3-receptor antagonist.[J]. Journal of Pharmacology and Experimental Therapeutics, 1998, 287 2: 658-666.