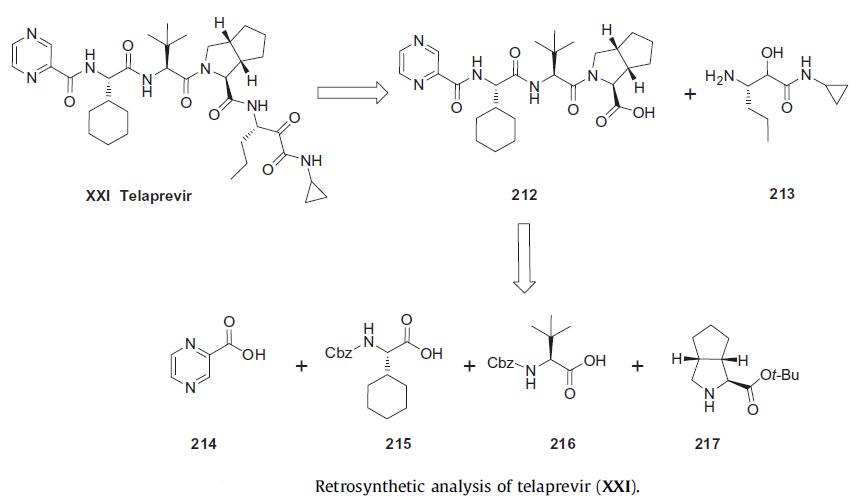
For preparation of bulk API, a convergent synthetic strategy
was utilized as described in the scheme. Retrosynthetically the
penultimate intermediate 212, which was coupled with amine
213 for the final step, was prepared by coupling bicyclic amine
217 with amino acids 216 and 215 and then with pyrazine acid
214.
In the early stages of development, the cyclopropyl amide fragment
213 was made using a MCR coupling sequence by reacting
aldehyde 218 with cyclopropyl isocyanide (219) and triflouroacetic
acid to give amide alcohol 220 in 85% yield. Removal
of the Cbz group was accomplished via hydrogenolysis to provide
key cyclopropyl amide alcohol 213 in 95% yield. While this route
was shorter in terms of steps, it was not amenable to large-scale
preparation due to difficulties associated with the handling of isocyanide
219.
Thus, for large-scale synthesis, the route depicted in Scheme 35
was utilized. Commercially available Cbz-protected amino acid
221 was converted to the corresponding Weinreb amide 222 using
CDI as the activating agent. This was followed by LAH reduction to
give aldehyde 218 in 73% yield from 221. Aldehyde 218 was reacted
with sodium cyanide under neutral to mildly basic conditions
allowing for easy workup of the cyanohydrin, which was
immediately hydrolyzed by refluxing in 4 N hydrochloric acid in
dioxane to deliver hydroxy acid HCl salt 223. Since the formation
of the acyloin resulted in removal of the Cbz protecting group, reinstallation
of this protecting group preceded conventional amide
bond formation through the intermediacy of the succinate ester
of 224. This provided the desired amide alcohol 220 in 56% yield
from 223. Hydrogenolysis of Cbz carbamate 220 then furnished
the requisite intermediate amine (213) in 73% yield.
The large-scale synthesis of bicyclic pyrrolidine 231 was accomplished
as described in the following scheme. Commercially available 3-azabicyclo[
3.3.0]nonane hydrochloride (225) was first protected as
the corresponding Boc carbamate 226 in 90% yield. Deprotonation
of the bicyclic pyrrolidine carbamate 226 with sec-BuLi and
sequential quench with bubbling carbon dioxide gas followed by
sodium hydrogensulfate resulted in racemic acid 227 in 80% yield.
Racemate 227 was resolved using (S)-tetrahydronapthalamine
(228) in ethyl acetate and isopropanol at 70¨C75C. This mixture
was allowed to cool down slowly to effect the crystallization of
the optically enriched chiral salt 229 in 83% yield with greater than
99.5% ee. This enatioenriched salt was free based with sodium
hydrogen sulfate and converted to t-butyl ester 230 using Boc
anhydride and DMAP. The secondary amine of 230 was liberated
using methane sulfonic acid at room temperature followed by salt formation with oxalic acid in isopropyl acetate to give oxalic acid
salt 231 in 81% yield over 3 steps.
With the synthesis of the key intermediates complete, sequential
coupling events were then executed to complete the synthesis
of teleprevir (following scheme). Fragment 217 was coupled with the
Cbz-protected valine (216) using EDCI and HOBt to give intermediate
232 in 87% yield. Similarly, after removal of the Cbz group of
232 via catalytic hydrogenolysis, the resulting amine was coupled
with cyclohexyl amino acid 215 to give dipeptide intermediate 233
in 89% yield over 2 steps. Sequential cleavage of the Cbz group in
233 followed by CDI-mediated coupling with commercially available
pyrazine acid 214 gave rise to the expected pyrazine amide
intermediate. Subsequent hydrolysis of the t-butyl ester through
the use of concentrated acid in DCM provided the key intermediate
tripeptidic acid (212) in 68% over 3 steps. The tripeptide 212 was
then coupled with cyclopropyl amide amine 213 using EDCI, HOBt
and N-methyl morpholine (NMM) to provide penultimate intermediate
alcohol 234 in 95% yield. Subjection of 234 to Dess¨CMartin
periodinane (DMP) oxidation in t-butanol and DCM furnished telaprevir
(XXI) in 85% yield.