Chemical Properties
Pale Yellow Solid
Uses
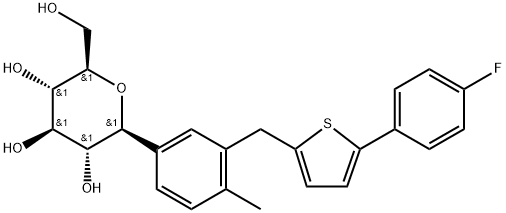
Canagliflozin is a sodium/glucose cotransporter 2 (SGLT2) inhibitor. Canagliflozin has been shown to dose dependently reduce calculated renal threshold for glucose excretion and increase urinary glucose excretion. Canagliflozin is a candidate for the treatment of type 2 diabetes and obesity.
Definition
ChEBI: A C-glycosyl compound that is used (in its hemihydrate form) for treatment of type II diabetes via inhibition of sodium-glucose transport protein subtype 2.
Description
In March 2013, the US FDA approved canagliflozin (JNJ-28431754;
TA-7284) for the treatment of type 2 diabetes in adults.
Canagliflozin is the first US-approved sodium-glucose co-transporter (SGLT) inhibitor for the treatment of type 2 diabetes. Inhibition of renal SGLT suppresses glucose reabsorption, which permits glucose excretion into urine and reduction of hyperglycemia. Canagliflozin was discovered through structural modifications of phlorizin, a known inhibitor of renal glucose reabsorption. Early modifications of OH groups on the glucose moiety were insufficient to adequately impair hydrolysis by intestinal β-glucosidase. Introduction of the C-glucoside moiety, as in the clinical candidate dapagliflozin, afforded sufficient resistance to hydrolysis. Finally, incorporation of the thiophene moiety in canagliflozin provided improved potency for hSGLT2 (exclusive to kidney), IC50=2.2 nM, while offering significant selectivity over hSGLT1 (in kidney and heart), IC50=910 nM. Hyperglycemic, high-fat (HF) diet fedmice (KKstrain) that received a single 3 mg/kg oral dose of canagliflozin had a 48% reduction in blood glucose levels after 6 h versus
vehicle-treatedmice. Noteworthy in themultistep synthesis of canagliflozin
is the stereoselective formation of the β-C-glucoside which is accomplished by coupling of the aryllithiumaglycone with 2,3,4,6-tetra-O-trimethylsilyl-β-Dgluconolactone followed by desilylation and stereoselective reduction with triethylsilane and boron trifluoride etherate.
Originator
Mitsubishi Tanabe
Pharma (Japan)
Synthesis Reference(s)
[1]Metil, D. S., Sonawane, S. P., Pachore, S. S., Mohammad, A., Dahanukar, V. H., McCormack, P. J., Reddy, C. V., & Bandichhor, R. (2017). Synthesis and Optimization of Canagliflozin by Employing Quality by Design (QbD) Principles. Organic Process Research & Development, 22(1), 27–39.
https://doi.org/10.1021/acs.oprd.7b00281
Clinical Use
Sodium-glucose co-transporter 2 inhibitor:
Treatment of type 2 diabetes
Synthesis
Canagliflozin is prepared through a six-step sequence that includes Friedel-Crafts acylation, Lewis acid mediated reductive elimination, n-BuLi mediated C-arylation and reductive demethoxylation, with the route proceeding via a purified open-ring intermediate[1]. Canagliflozin is also obtained by β-selective C-arylation of silyl-protected 1,6-anhydroglucose with an arylalane, or by C-arylation of 1,6-anhydroglucose modified with diisobutylaluminium hydride using an arylalane[1].

Drug interactions
Potentially hazardous interactions with other drugs
Antibacterials: concentration reduced by rifampicin.
Lipid-regulating drugs: avoid canagliflozin for 1 hour
before or 4-6 hours after bile acid sequestrants.
Metabolism
O-glucuronidation is the major metabolic elimination
pathway mainly by UGT1A9 and UGT2B4 to two
inactive metabolites. CYP3A4-mediated (oxidative)
metabolism of canagliflozin is minimal (approximately
7%) in humans.